Launching a product in the European Union is not simply a matter of sending a finished sample to a laboratory.
By the time laboratory testing begins, many important decisions should already be clear.
What is the product?
Which EU rules apply?
Are the ingredients permitted?
What claims are planned?
Which documents are required?
And, most importantly, what do you actually need to prove?
Preparing these questions early can make laboratory testing more targeted, reduce unnecessary analysis and prevent regulatory problems from appearing late in the launch process.
EU market entry starts before laboratory testing
A common approach is:
Develop the product → test it → check compliance → launch
That sequence can create problems.
A better approach is:
Classify the product → identify requirements → assess risks → define evidence → test → document → launch
The difference is important.
Laboratory testing should answer questions that have already been identified through the regulatory and product assessment.
Otherwise, companies can end up with a comprehensive laboratory report that still does not demonstrate EU compliance.
Step 1: Define what your product actually is
Before choosing any laboratory test, determine the product category.
Is it:
- a food?
- a food supplement?
- a cosmetic?
- a pharmaceutical product?
- a medical device?
- a food-contact material?
- another consumer product?
The classification determines which legislation, standards and regulatory requirements apply.
For example, food supplements are regulated as foods under the EU framework, while cosmetics are subject to Regulation (EC) No 1223/2009. Medical devices follow a separate regulatory framework under Regulation (EU) 2017/745.
This first classification therefore influences almost every later decision.
Why classification matters
Imagine a company has developed a product containing a botanical extract.
The company wants to sell it as a wellness product.
That description alone is not enough.
Depending on the composition, presentation and intended use, different regulatory questions may arise.
The product could be assessed as:
- a food
- a food supplement
- a cosmetic
- or potentially another regulated product
The testing programme would then look very different.
So before asking:
“What should we test?”
ask:
“What regulatory category does our product belong to?”
Step 2: Define the intended market
“EU market” sounds like one market.
Regulatory requirements are more nuanced.
EU legislation can be harmonised at Union level, while certain areas also involve national provisions or national enforcement practices.
You should therefore define:
- which EU countries you plan to enter
- whether the product will be sold online
- whether it will be distributed through retailers
- whether it will be imported
- who will act as the Responsible Person or importer where required
This information helps determine the regulatory pathway.
Step 3: Review the formulation
Once the product category is clear, examine the complete formulation.
Create a list of:
- active ingredients
- functional ingredients
- excipients
- preservatives
- fragrances
- colours
- botanical extracts
- vitamins
- minerals
- processing aids
Then ask:
Is every ingredient permitted for this product category and intended use?
This should happen before expensive testing begins.
There is little value in fully testing a formulation if a key ingredient creates a fundamental regulatory problem.
Step 4: Check ingredient restrictions
An ingredient can be legally available and still be restricted in a particular product.
For cosmetics, for example, the EU Cosmetics Regulation contains prohibited substances in Annex II and restricted substances in Annex III, while preservatives, colourants and UV filters are addressed through additional annexes. (eur-lex.europa.eu)
The same principle applies across other regulated product categories.
Therefore, ingredient screening should consider:
- identity
- concentration
- purity
- product category
- intended use
- target population
- applicable restrictions
A simple ingredient name check is not enough.
Step 5: Check whether an ingredient has special regulatory status
Some ingredients require additional attention.
For food and supplements, this can include potential Novel Food status.
Under Regulation (EU) 2015/2283, foods not consumed to a significant degree in the EU before 15 May 1997 can fall within the Novel Food framework and may require authorisation before being placed on the EU market. (food.ec.europa.eu)
Checking this early is critical.
A laboratory can analyse an ingredient.
It cannot make an unauthorised Novel Food authorised.
Step 6: Review the claims
Claims should be assessed before testing, not after.
Ask:
What do we want the product to say?
For example:
- high protein
- source of fibre
- supports skin health
- moisturising
- antioxidant
- clinically tested
- microbiologically tested
Each type of claim can create different evidence requirements.
For foods, EU Regulation (EC) No 1924/2006 governs nutrition and health claims, while authorised health claims are listed in the EU Register. (food.ec.europa.eu)
The claim strategy can therefore influence the analytical strategy.
Step 7: Determine what needs to be proven
Now turn regulatory requirements into evidence requirements.
For example:
Requirement
The product must contain a defined amount of an active ingredient.
Evidence
Quantitative analytical testing.
Or:
Requirement
The product must remain stable throughout its proposed shelf life.
Evidence
A suitable stability programme.
Or:
Requirement
The packaging must be suitable for food contact.
Evidence
Appropriate migration and supporting documentation.
This creates a direct connection between:
requirement → evidence → test
That connection is one of the most important parts of efficient market preparation.
Step 8: Assess the raw materials
Before testing the finished product, review the raw materials.
For each critical ingredient, consider:
- supplier
- origin
- specification
- manufacturing process
- Certificate of Analysis
- contaminant risks
- microbiological risks
- relevant impurities
- regulatory status
A strong raw-material assessment can prevent unnecessary finished-product testing.
It can also identify risks before formulation is finalised.
A supplier COA is useful, but not enough
A Certificate of Analysis can provide valuable information about a raw material.
However, it does not automatically demonstrate EU compliance of the finished product.
The COA may only cover selected parameters.
The finished product may have additional requirements.
Therefore, supplier documentation should be reviewed as part of the evidence package rather than treated as a complete compliance assessment.
Step 9: Identify the product’s main risks
Now create a risk map.
Depending on the product, this may include:
Chemical risks
- heavy metals
- pesticides
- mycotoxins
- residual solvents
- impurities
- degradation products
Microbiological risks
- pathogens
- total microbial count
- yeasts and moulds
Physical risks
- particle size
- viscosity
- dissolution
- homogeneity
Stability risks
- oxidation
- moisture
- pH changes
- active degradation
Packaging risks
- migration
- moisture transfer
- oxygen exposure
- compatibility
The testing plan should then focus on the risks that actually apply.
Step 10: Build the testing strategy
Only now should you start defining the laboratory programme.
A risk-based plan might include:
| Question | Possible evidence |
|---|---|
| Is the ingredient correctly identified? | Identity testing |
| Does the product contain the expected amount? | Quantitative analysis |
| Are contaminants controlled? | Contaminant analysis |
| Is the product microbiologically acceptable? | Microbiological testing |
| Will it remain within specification? | Stability testing |
| Is packaging suitable? | Migration/compatibility testing |
| Can a claim be supported? | Analytical or scientific evidence |
The exact tests depend on the product.
The objective is not to order every available analysis.
It is to generate the evidence needed for the EU market.
Step 11: Check the analytical methods
Once you know what needs to be tested, consider how it should be measured.
The method should be appropriate for:
- the analyte
- the matrix
- the expected concentration
- the regulatory limit
- the intended decision
Important characteristics can include:
- specificity
- accuracy
- precision
- sensitivity
- detection limit
- quantification limit
For pharmaceutical products, ICH Q2(R2) provides a framework for validation of analytical procedures and their performance characteristics. (ema.europa.eu)
The principle is broader than pharmaceuticals:
The analytical method should be suitable for the regulatory question it is being used to answer.
Step 12: Define acceptance criteria before testing
Do not wait for the laboratory result before deciding whether it is acceptable.
Define the relevant criterion first.
That could be:
- a legal maximum
- a minimum content
- an internal specification
- a microbiological criterion
- a stability specification
For example:
Target: 95–105%
Laboratory result: 102%
Conclusion: Meets the defined specification.
This makes the decision process transparent.
Step 13: Consider the finished product
Raw-material testing and finished-product testing answer different questions.
A raw material may meet its specification.
The finished product can still behave differently because of:
- processing
- interactions
- degradation
- contamination
- concentration
- packaging
Therefore, the final product needs to be assessed against its own specifications.
Step 14: Consider stability early
Stability testing should not be left until the final weeks before launch.
The results can influence:
- shelf life
- packaging
- storage conditions
- formulation
- product specifications
If the product is unstable, the company needs enough time to make changes.
Starting early therefore reduces the risk of a late-stage reformulation.
Step 15: Review the packaging
Packaging can influence both compliance and product stability.
Depending on the product, consider:
- migration
- compatibility
- moisture protection
- oxygen protection
- light protection
- mechanical protection
For food-contact materials, EU rules establish general safety requirements and specific measures for different material groups. (food.ec.europa.eu)
The package should therefore be considered part of the product system.
Step 16: Build the documentation alongside testing
Do not wait until all laboratory reports are finished before organising the technical documentation.
Depending on the product, this can include:
- formulation
- raw-material specifications
- supplier documentation
- laboratory reports
- stability data
- safety assessments
- product specifications
- claims evidence
- labelling
- packaging information
- manufacturing information
Keeping this documentation aligned from the beginning makes the final compliance review much easier.
Different products need different EU entry strategies
Cosmetics
Key areas can include:
- formulation compliance
- prohibited and restricted ingredients
- safety assessment
- Product Information File
- labelling
- claims
- microbiological quality
- stability
The EU Cosmetics Regulation requires a safety assessment before a cosmetic product is placed on the market and establishes requirements for the Product Information File. (eur-lex.europa.eu)
Food and supplements
Important areas can include:
- ingredient status
- contaminants
- microbiology
- composition
- nutrition information
- claims
- Novel Food status
- stability
- packaging
Food supplements are regulated under EU food law, including Directive 2002/46/EC. (eur-lex.europa.eu)
Pharmaceuticals
The requirements are more extensive and formalised.
They can involve:
- pharmaceutical quality
- specifications
- analytical methods
- stability
- manufacturing
- impurities
- validation
- regulatory submissions
EMA quality guidelines cover areas including specifications, analytical procedures, analytical validation and stability. (ema.europa.eu)
Medical devices
Medical-device market entry involves its own regulatory pathway under the EU Medical Device Regulation.
Depending on the device, manufacturers need to consider:
- classification
- risk management
- biological safety
- chemical characterisation
- microbiological safety
- technical documentation
- conformity assessment
- clinical evidence where applicable
Regulation (EU) 2017/745 establishes the EU framework for medical devices. (eur-lex.europa.eu)
Again, laboratory testing is only one part of the process.
Why testing too early can also be inefficient
There is another side to this.
Starting laboratory testing before the regulatory questions are clear can create unnecessary work.
For example, a company might commission:
- a broad contaminant panel
- multiple stability studies
- unnecessary active-content testing
- tests for parameters that are not relevant to the final product
Later, the formulation changes.
Now some of the results may no longer represent the product.
That is why regulatory scoping should come before the final testing plan.
The ideal sequence
A practical EU market-entry workflow looks like this:
1. Product classification
↓
2. Target market
↓
3. Ingredient review
↓
4. Regulatory screening
↓
5. Claims review
↓
6. Risk assessment
↓
7. Testing strategy
↓
8. Laboratory testing
↓
9. Stability and supporting studies
↓
10. Regulatory documentation
↓
11. Final compliance review
↓
12. Market entry
The exact sequence can vary.
However, the principle remains:
Know what you need to prove before you start generating evidence.
What should you have ready before contacting a laboratory?
A laboratory can work much more efficiently when it receives sufficient product information.
Ideally, prepare:
- product category
- target market
- complete formula
- ingredient concentrations
- raw-material specifications
- intended use
- target population
- proposed claims
- packaging information
- manufacturing process
- proposed shelf life
- existing test data
This information allows the testing programme to be tailored to the actual product.
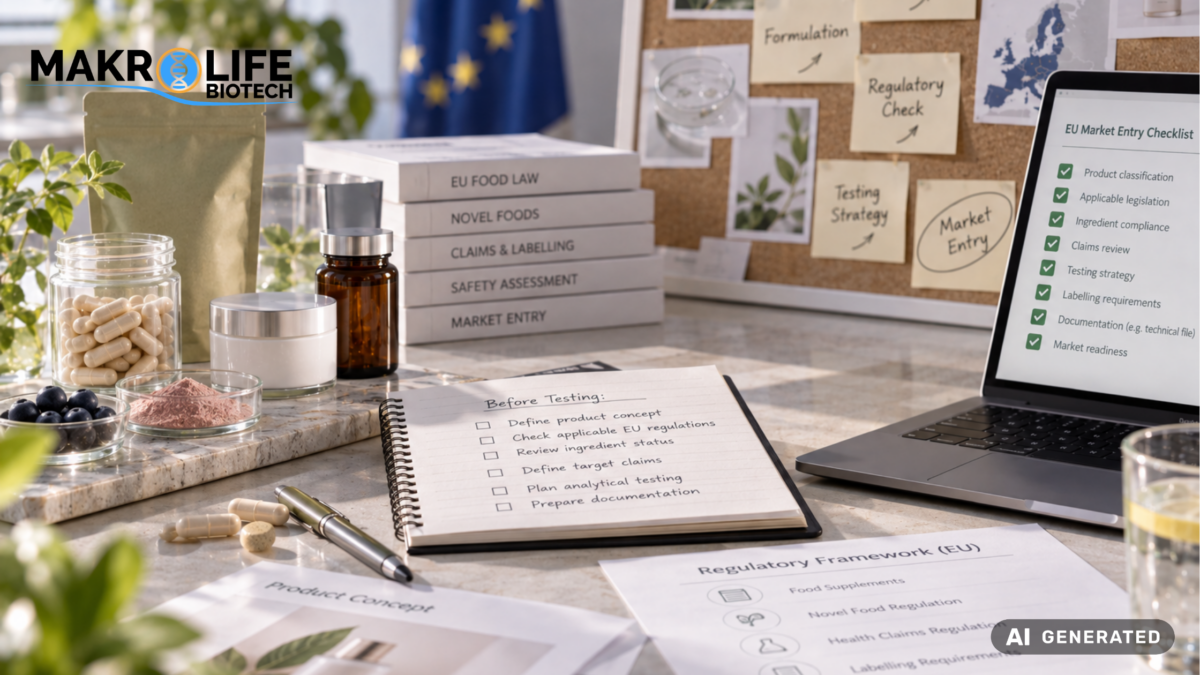
A simple pre-testing checklist
Before commissioning your first laboratory test, ask:
Product
What exactly are we placing on the EU market?
Regulation
Which legislation applies?
Ingredients
Are all ingredients permitted for the intended use?
Claims
What are we planning to communicate?
Risks
Which chemical, microbiological or physical risks are relevant?
Specifications
What acceptance criteria apply?
Testing
Which tests generate the evidence we actually need?
Documentation
Where will the results be used?
Changes
What happens if the product or supplier changes later?
If these questions have clear answers, laboratory testing becomes much more focused.
The biggest mistake: testing before knowing why
A company can spend thousands on laboratory analysis and still have an incomplete EU compliance package.
That happens when testing is treated as the starting point.
It is not.
The starting point is the regulatory question.
For example:
Requirement:
Contaminant must remain below the applicable limit.
Risk:
Ingredient may contain the contaminant.
Evidence:
Appropriate analytical test.
Decision:
Result meets or exceeds the applicable criterion.
That is a complete evidence chain.
What happens if the first test fails?
A failed test is easier to manage when the regulatory strategy was defined beforehand.
You can ask:
- Is the result reliable?
- Is the method appropriate?
- Does the batch actually fail?
- Can the formulation be adjusted?
- Can the raw material be replaced?
- Does the specification need review?
- Is additional evidence required?
Without a defined regulatory framework, a failed result can quickly turn into a confusing development problem.
EU market entry is an evidence-building process
The goal is not simply to collect laboratory reports.
It is to build a coherent body of evidence showing that the product is:
correctly classified
appropriately formulated
safe for its intended use
within applicable limits
supported by appropriate testing
correctly labelled
and properly documented.
Laboratory testing is an important part of that process.
It is not the entire process.
How Makrolife can support the process
This is where Makrolife can become involved before the first laboratory test.
The starting point can be a regulatory and product assessment.
From there, the testing strategy can be aligned with the actual requirements of the product.
Depending on the product category, this can include:
- regulatory assessment
- ingredient review
- risk assessment
- analytical testing
- stability studies
- safety assessment
- conformity assessment
- technical documentation
- EU market-entry support
The advantage is that the testing programme is built around the regulatory objective, rather than being selected independently from it.
The bottom line
EU market entry should not start with:
“Which laboratory tests do we need?”
It should start with:
“What do we need to demonstrate before this product can be placed on the EU market?”
Once that question is answered, the rest becomes much clearer.
First, classify the product.
Then identify the applicable requirements.
Review the ingredients and claims.
Assess the relevant risks.
Define the evidence you need.
Only then build the laboratory testing programme.
That approach can reduce unnecessary testing, identify problems earlier and create a much stronger compliance file.
The best time to plan EU compliance is before the first laboratory test, not after the final result comes back.